Welcome to PreCSenM!
The Predictive Cellular Senescence Model (PreCSenM) is a machine learning–based tool designed for senescence quantification. It enables users to quantify cellular senescence, identify senescence-associated features, and explore senescence landscapes across senescent cell line evaluation cohorts, lung adenocarcinoma (LUAD), pan-cancer types from TCGA, and single-cell RNA sequencing datasets.
Senescence Estimation
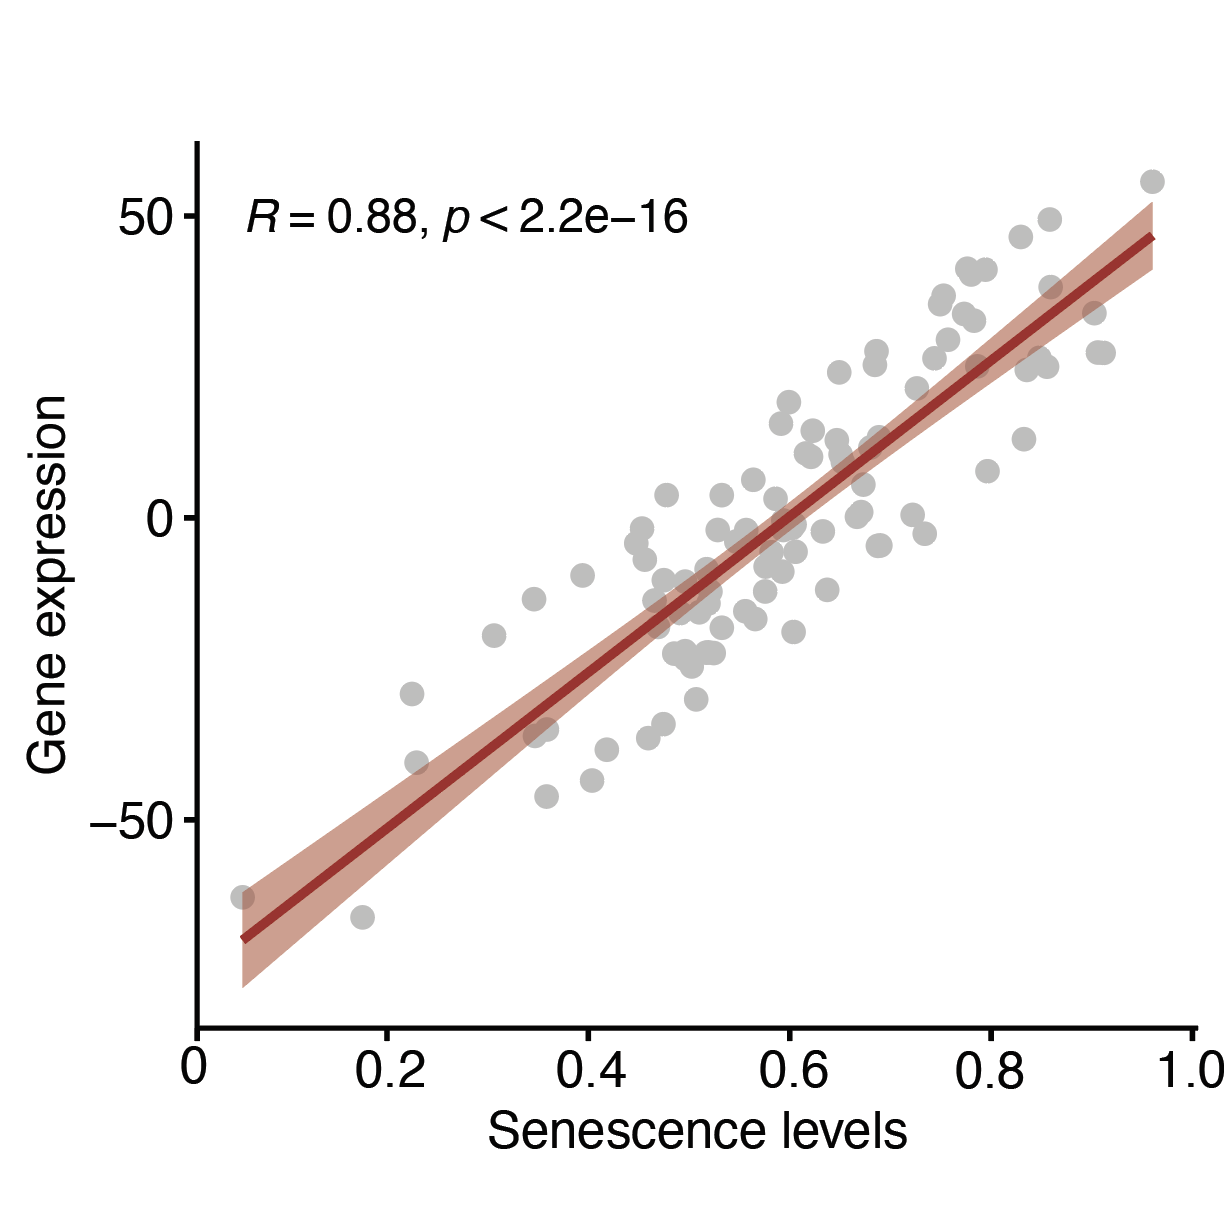
Estimation of cellular senescence (CS) levels and senescence-associated features in user-customized files
Senescence Exploration
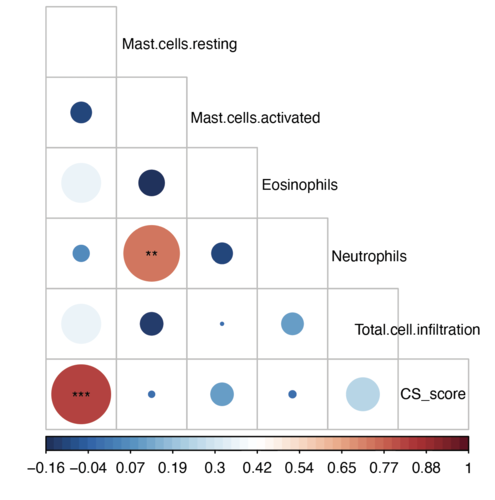
Explore senescence-associated characteristics in senescent cell line cohorts, lung adenocarcinoma (LUAD), TCGA cancer datasets, and single-cell RNA-seq data Senescent Cell Line Analysis
Predict senescence states in both normal and cancer-related transcriptomic datasets
Lifei Ma, Huiyang Li, ... Gong-Hong Wei, Xiaoman Wang, Hou-Zao Chen. Machine learning-based definition of cellular senescence reveals pro-senescence potential implications in lung adenocarcinoma. Genome Medicine. 2026.
Senescence Quantification and Identification of Senescence-Associated Characteristics
Explore senescence-associated features in diverse public datasets.
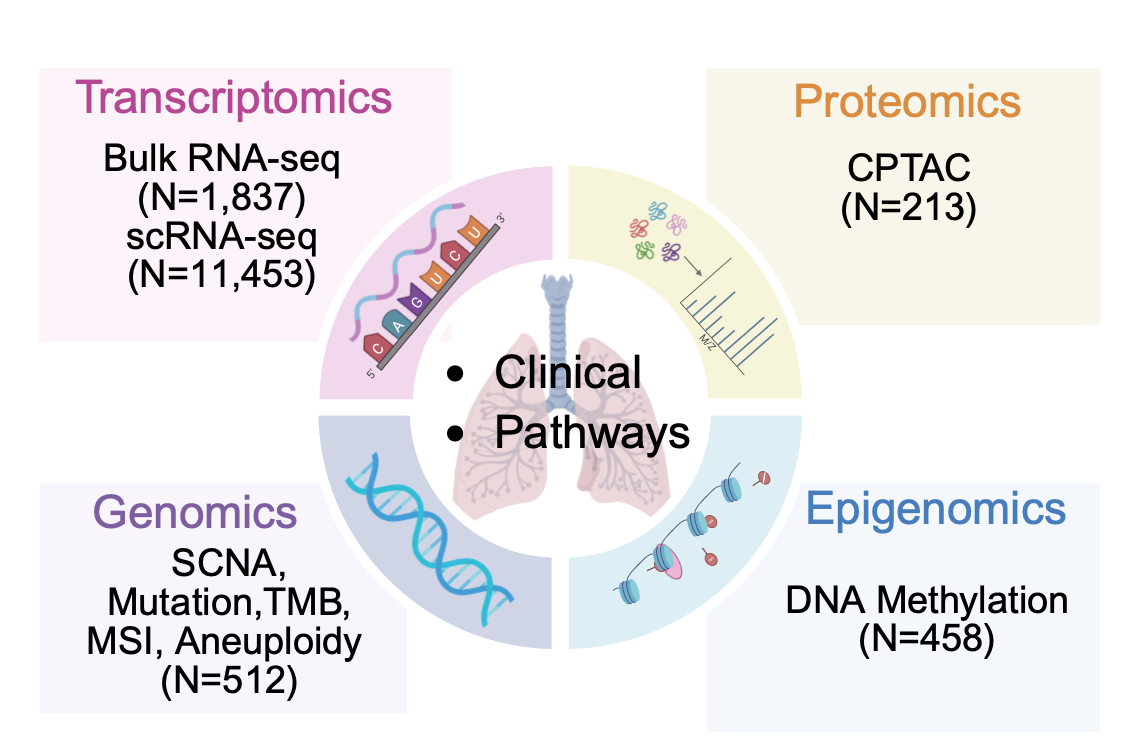
In LUAD, our analyses explored the associations between senescence levels and patient survival, genomic alterations, DNA methylation changes, proteogenomic profiles, and pathway enrichment.
The senescent cell line prediction module allows users to investigate senescence states across diverse data types, cell types, and senescence induction methods.
AUC plot showcasing selected data types, cell types, and senescence induction methods based on the CS scores.
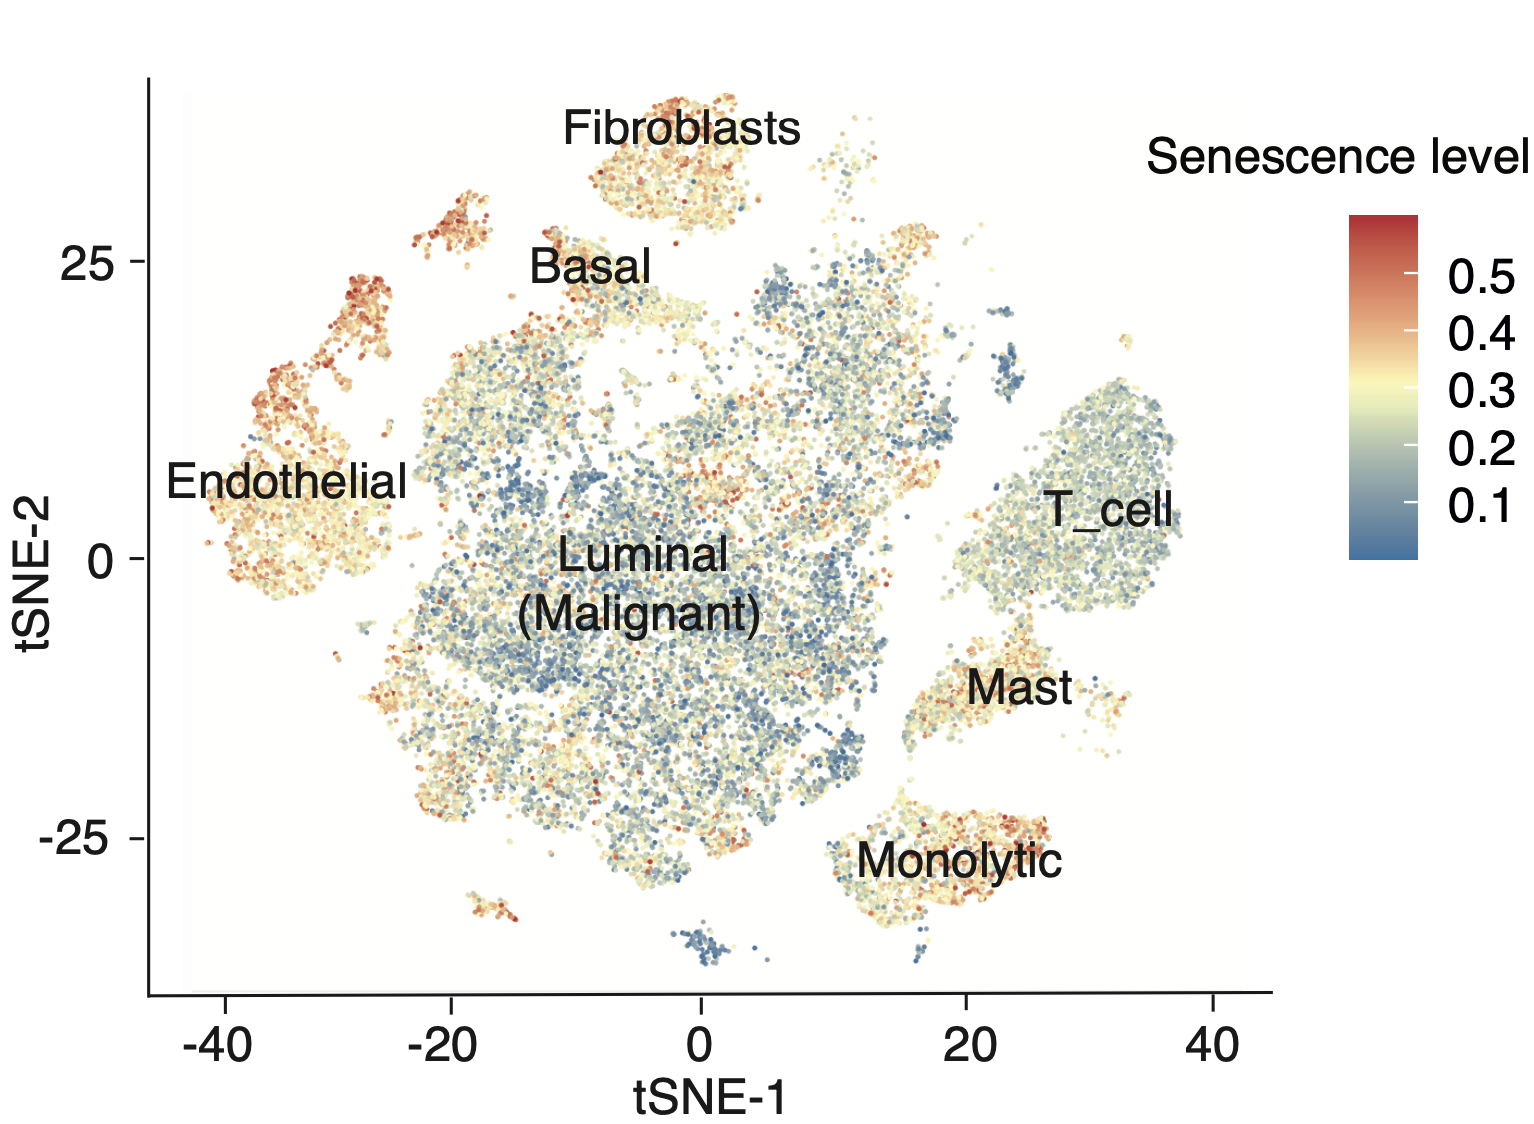
Single-cell analysis module allows users to explore senescence heterogeneity in diverse tumor microenvironment.
UMAP plots show major cell types in selected dataset. Box plot compares the difference of senescence levels (CS scores) across major cell types.
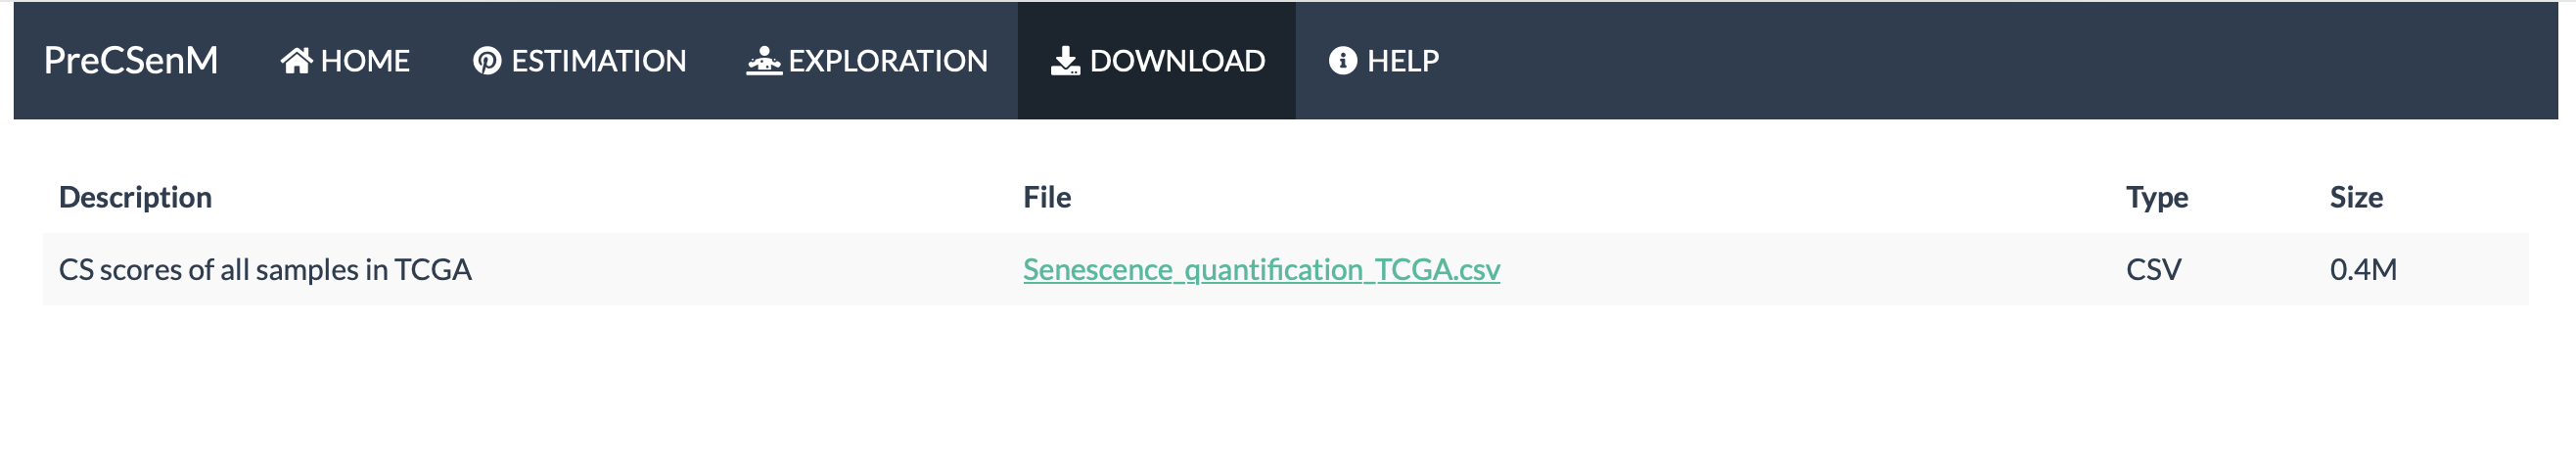
| Description | File | Type | Size |
|---|---|---|---|
| CS scores of all samples in TCGA | Senescence_quantification_TCGA.csv | CSV | 0.4M |
Introduction
Predictive Cellular Senescence Model(PreCSenM) is a comprehensive resource for systematical quantification of cellular senescence across cancer types. PreCSenM provides senescence quantification and identification of senescence-associated characteristics, and allow users to explore senescence across TCGA cancer types, immunotherapy cohorts and single-cell datasets.
Establish Predictive Cellular Senescence Model (PreCSenM)
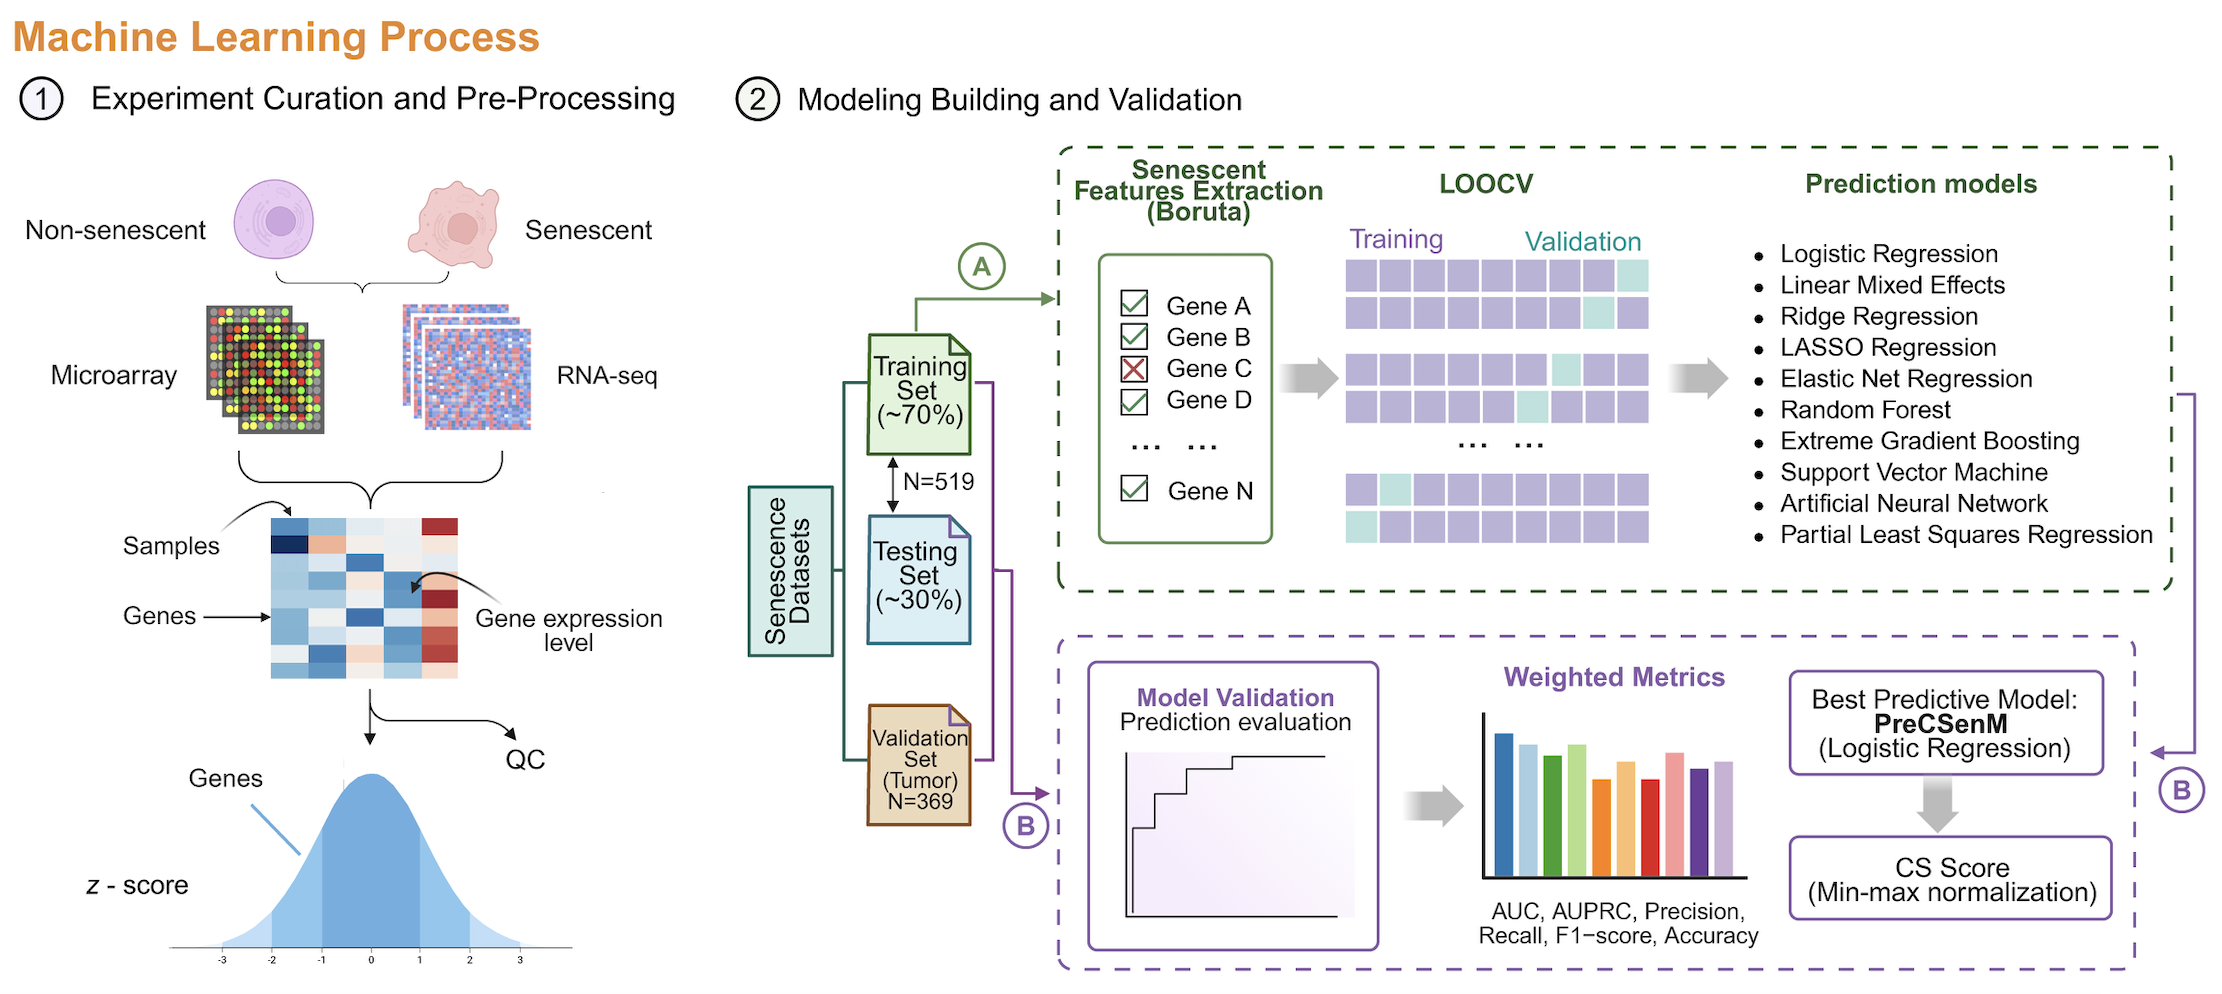
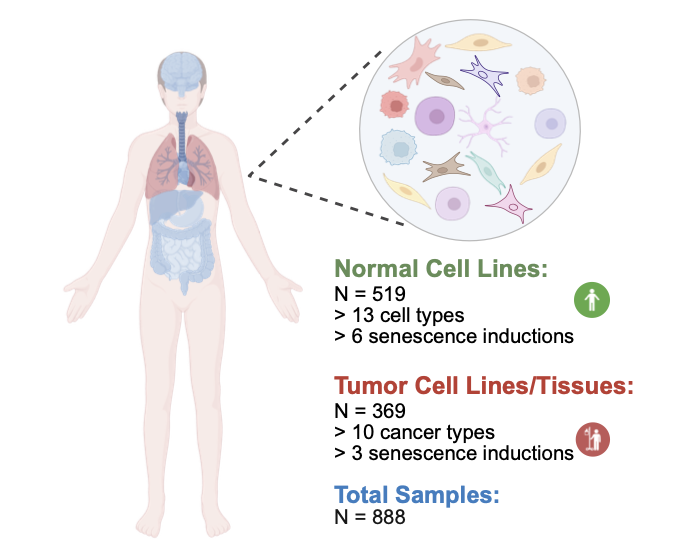
We established PreCSenM in three steps. First, we conducted experimental curation and pre-processing, including finding and curating 888 publicly available experimental series in RNA-seq and microarray data of normal cell lines and tumor cell lines or samples, performing quality control metrics, and computing z-scores per experiment. Second, we extracted senescent feature using the Boruta machine learning model to identify key senescence-associated genes by distinguishing between senescent and non-senescent responses; and model building and validation, involving the application of ten machine learning models and evaluating model performance using multiple metrics.
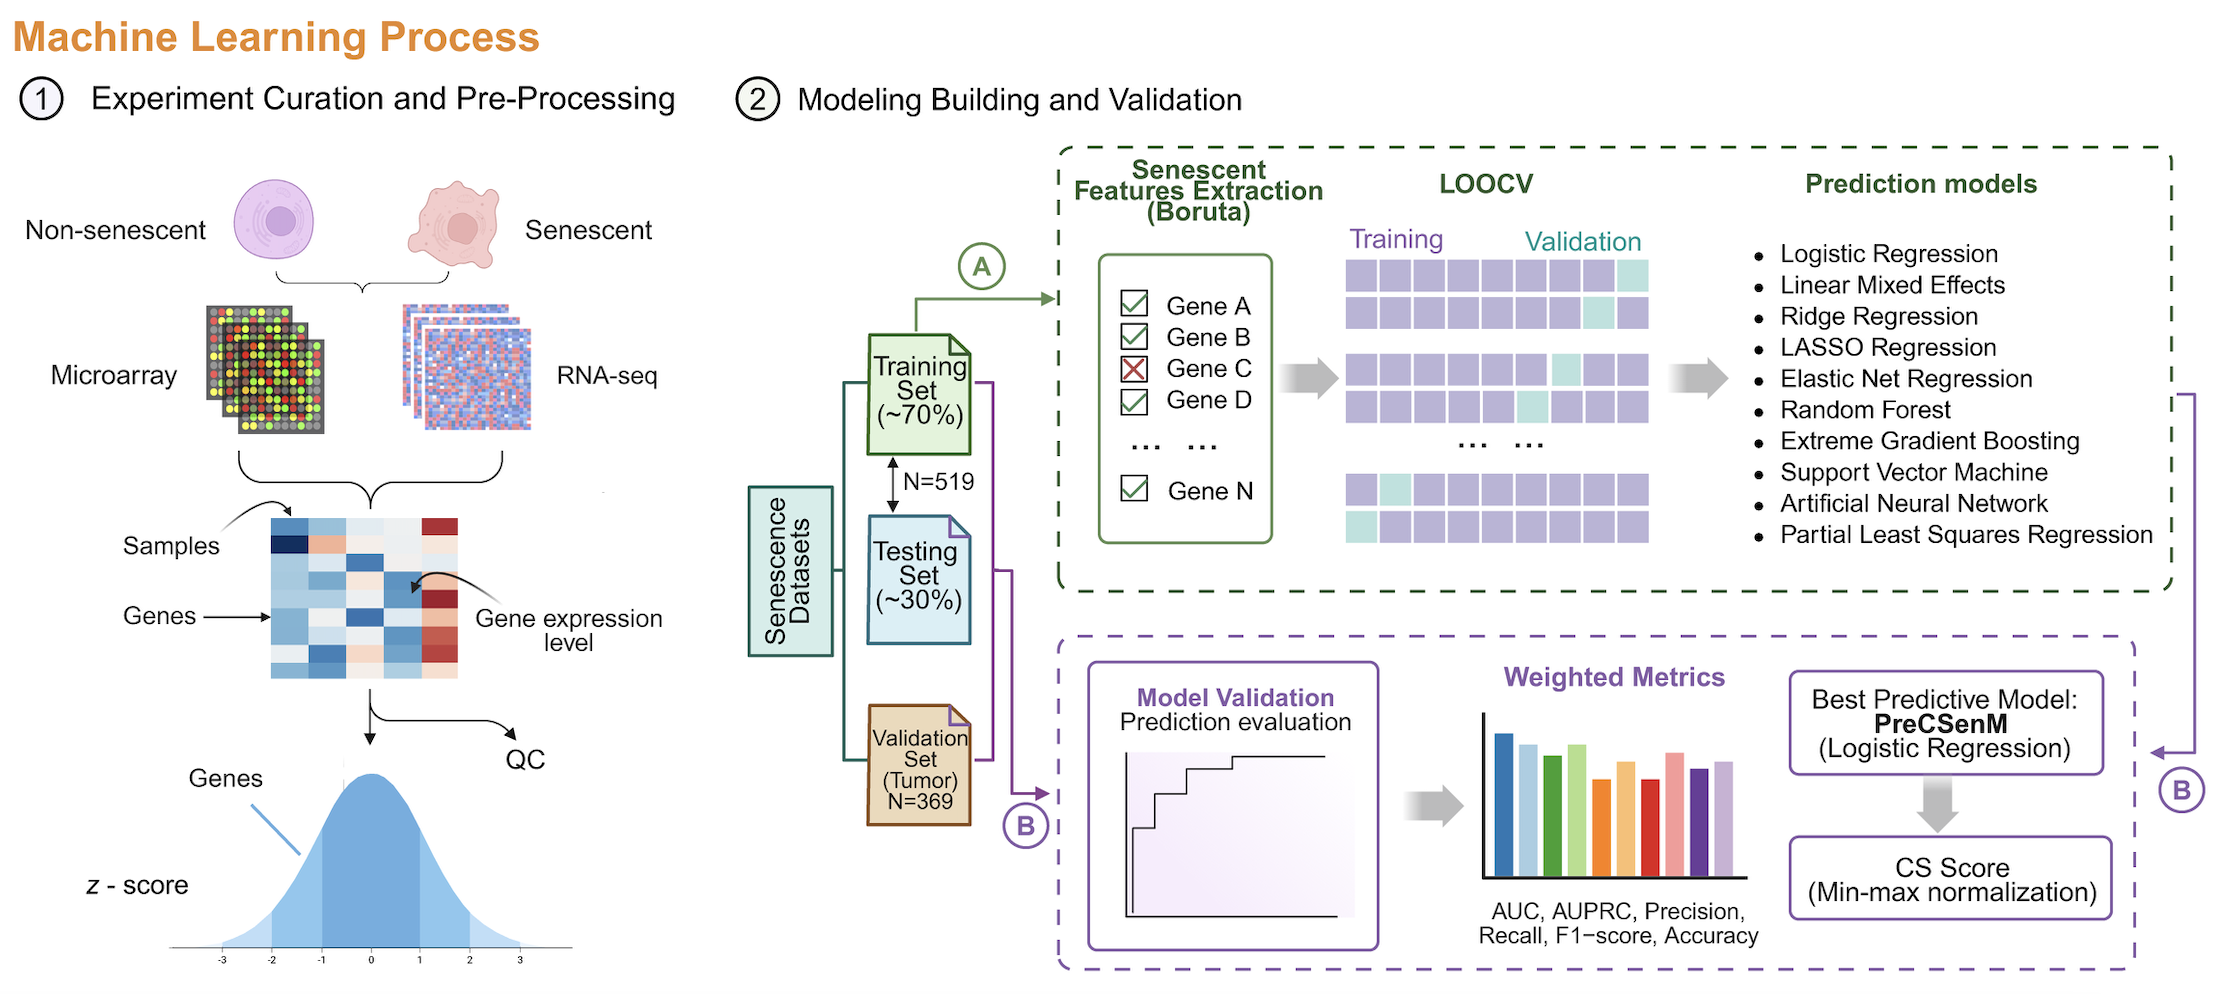
Workflow of PreCSenM
Function of PreCSenM
*Estimation of senescence level in user-customized file
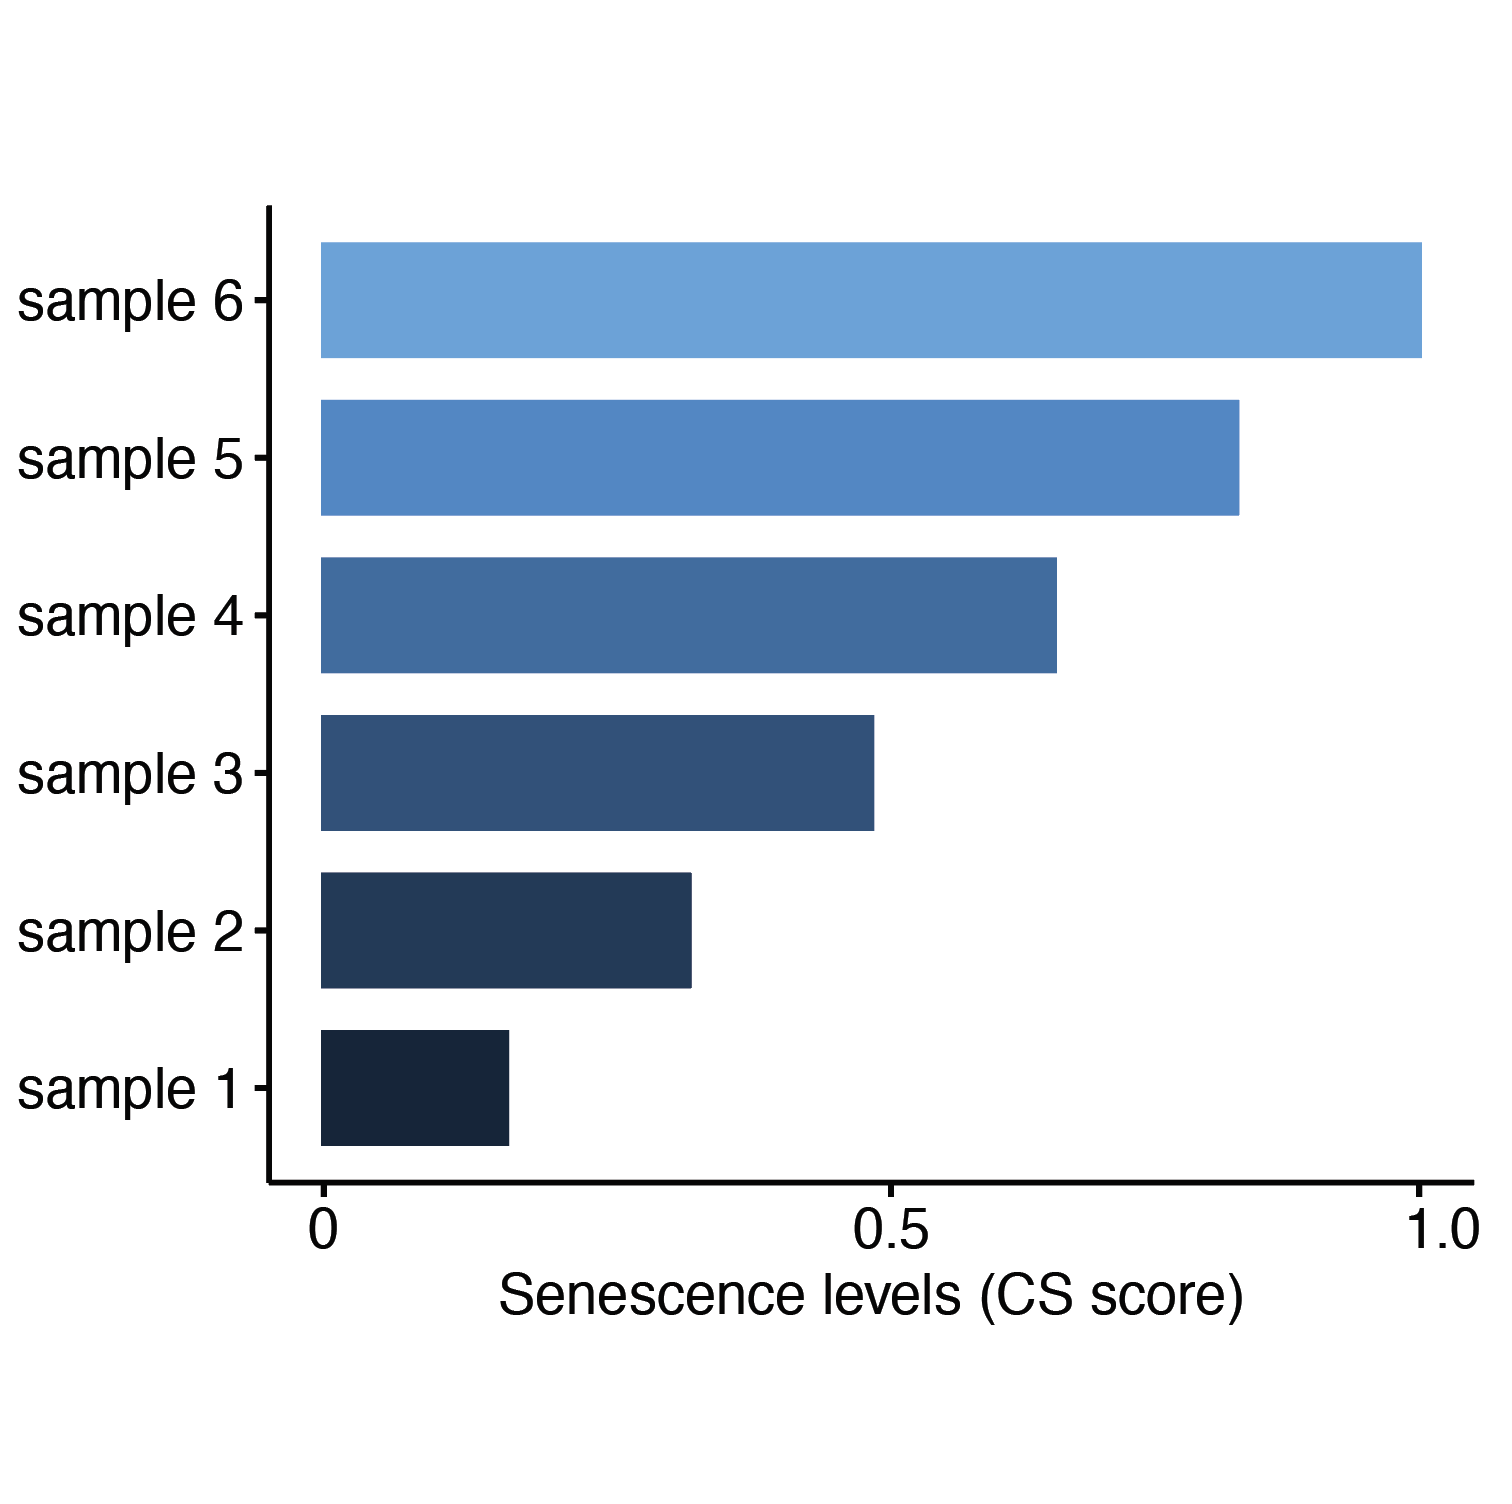
Users are allowed to upload gene expression matrix and the website will generate high-quality figures to estimate senescence level across samples, and list senescence-related genes and cell types.
*Exploration of senescence landscape in pan-cancer levels
We also investigated the associations between senescence levels and various biological and clinical features across senescent cell line evaluation cohorts, lung adenocarcinoma (LUAD), and pan-cancer types from TCGA. In LUAD, our analyses included predictions of genomic alterations, patient survival, and changes in DNA methylation and proteogenomic profiles. Additionally, we applied PreCSenM to publicly available single-cell transcriptomic datasets to compare senescence levels across major cell types.
Estimation
In the senescence estimation section, users are allowed to upload gene expression matrix (csv file) and specify the analysis of interest.
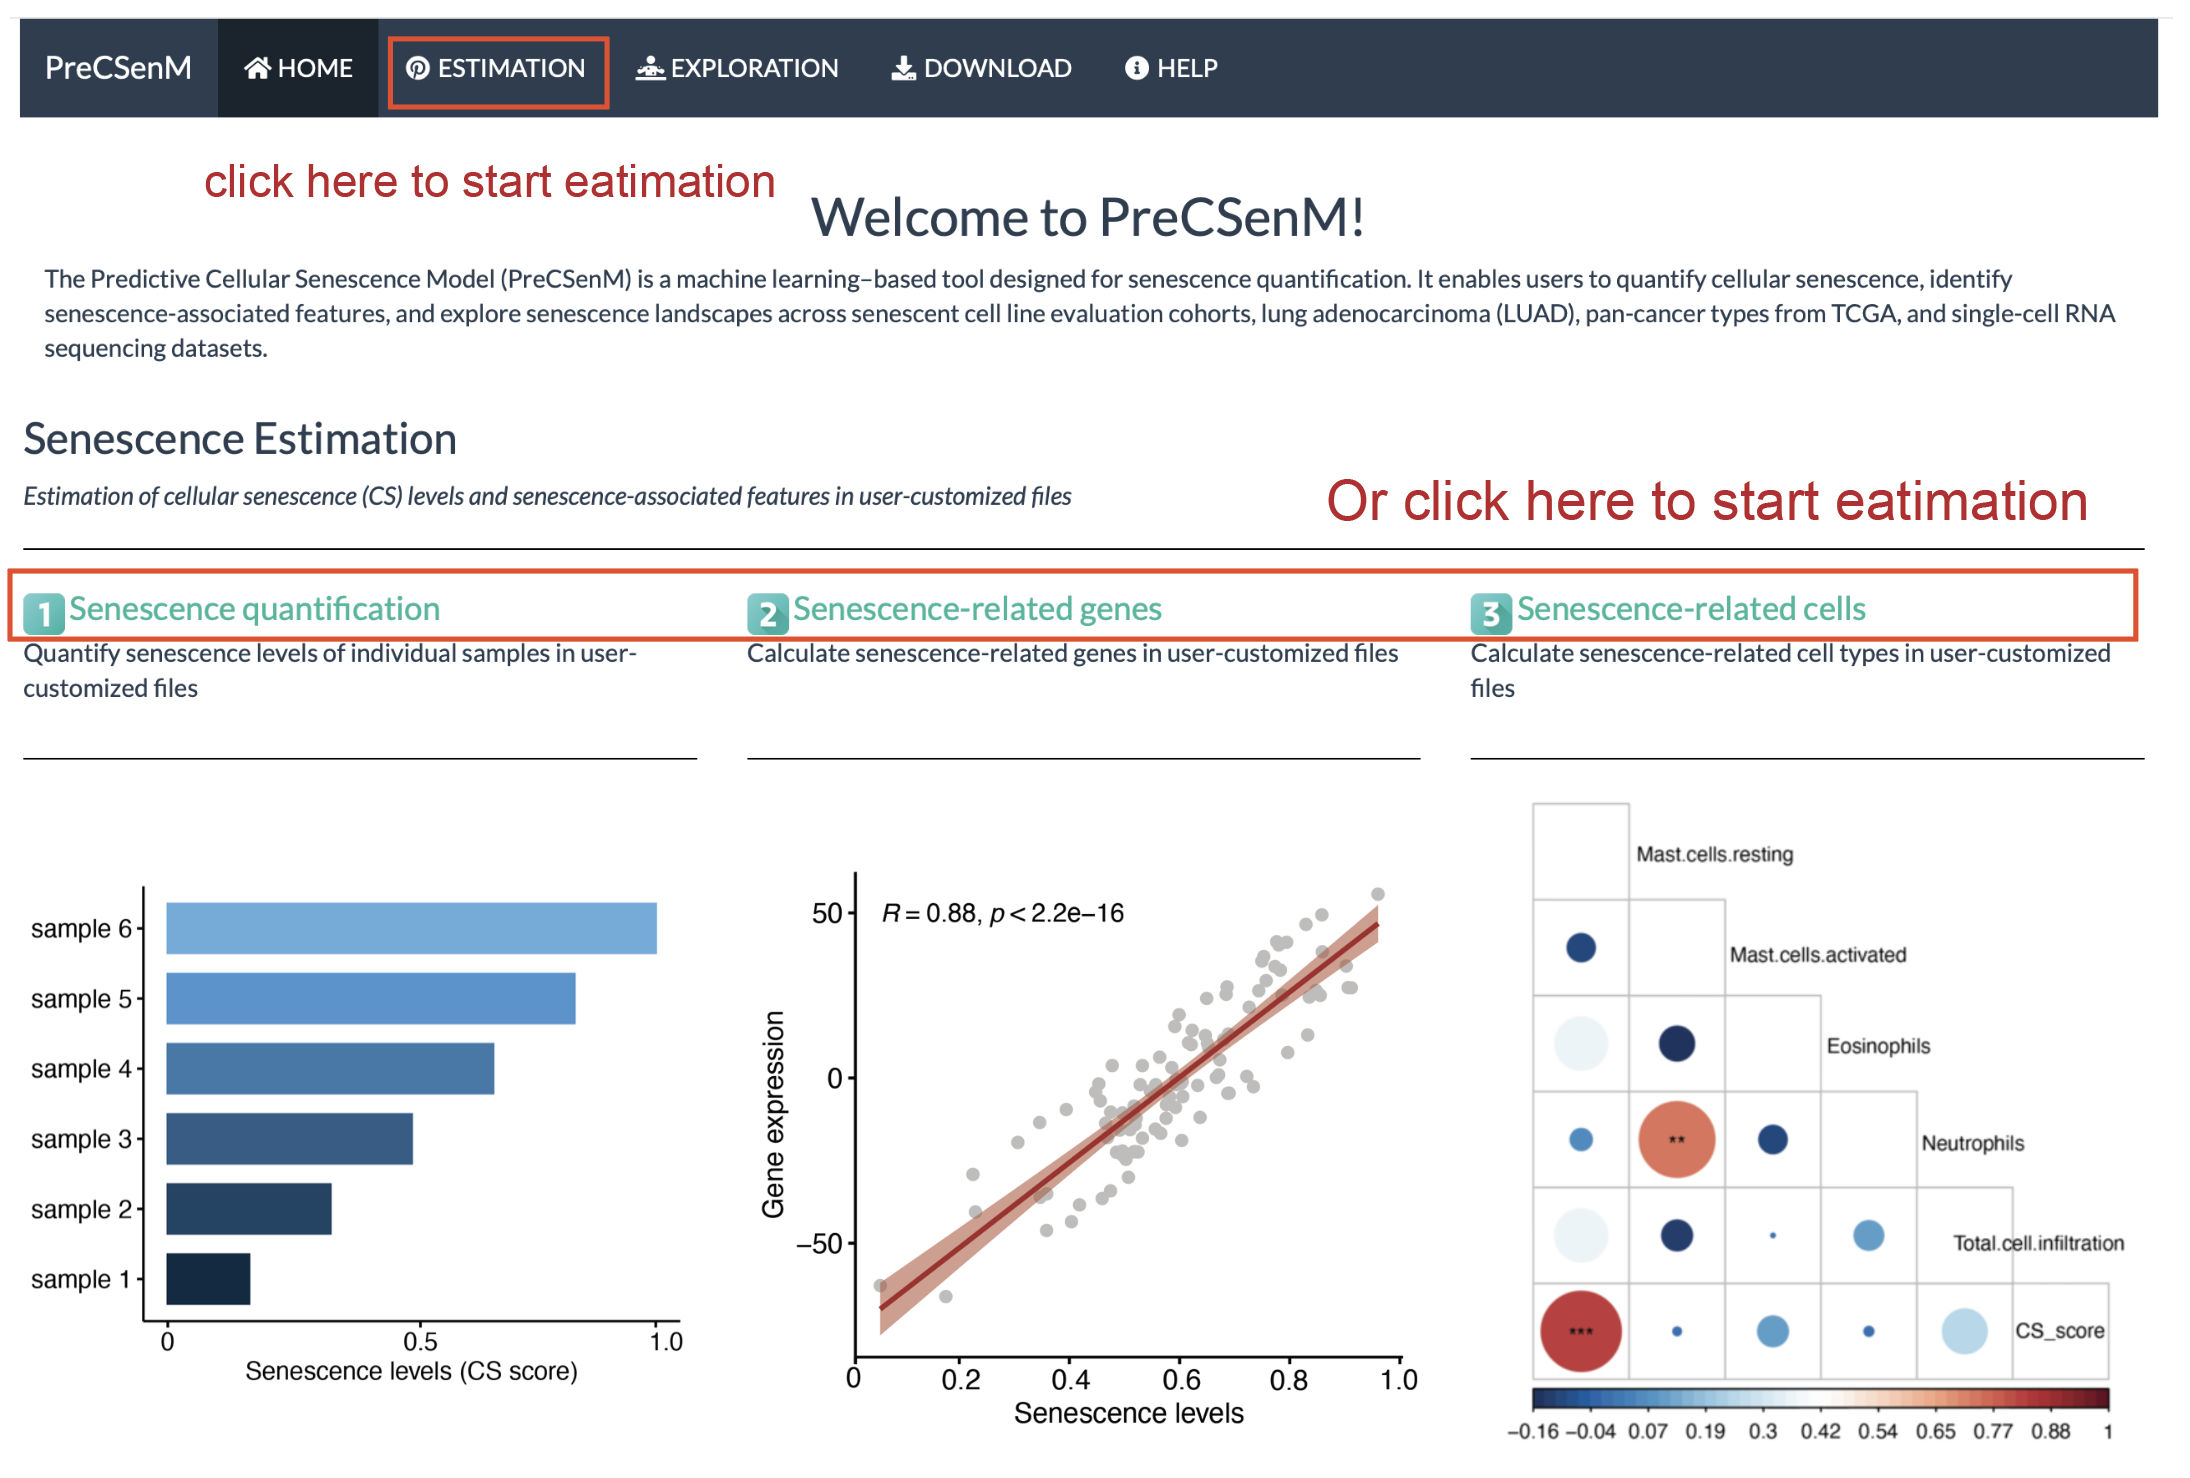
Senescence quantification and Identify senescence-associated cells may cost several minutes.
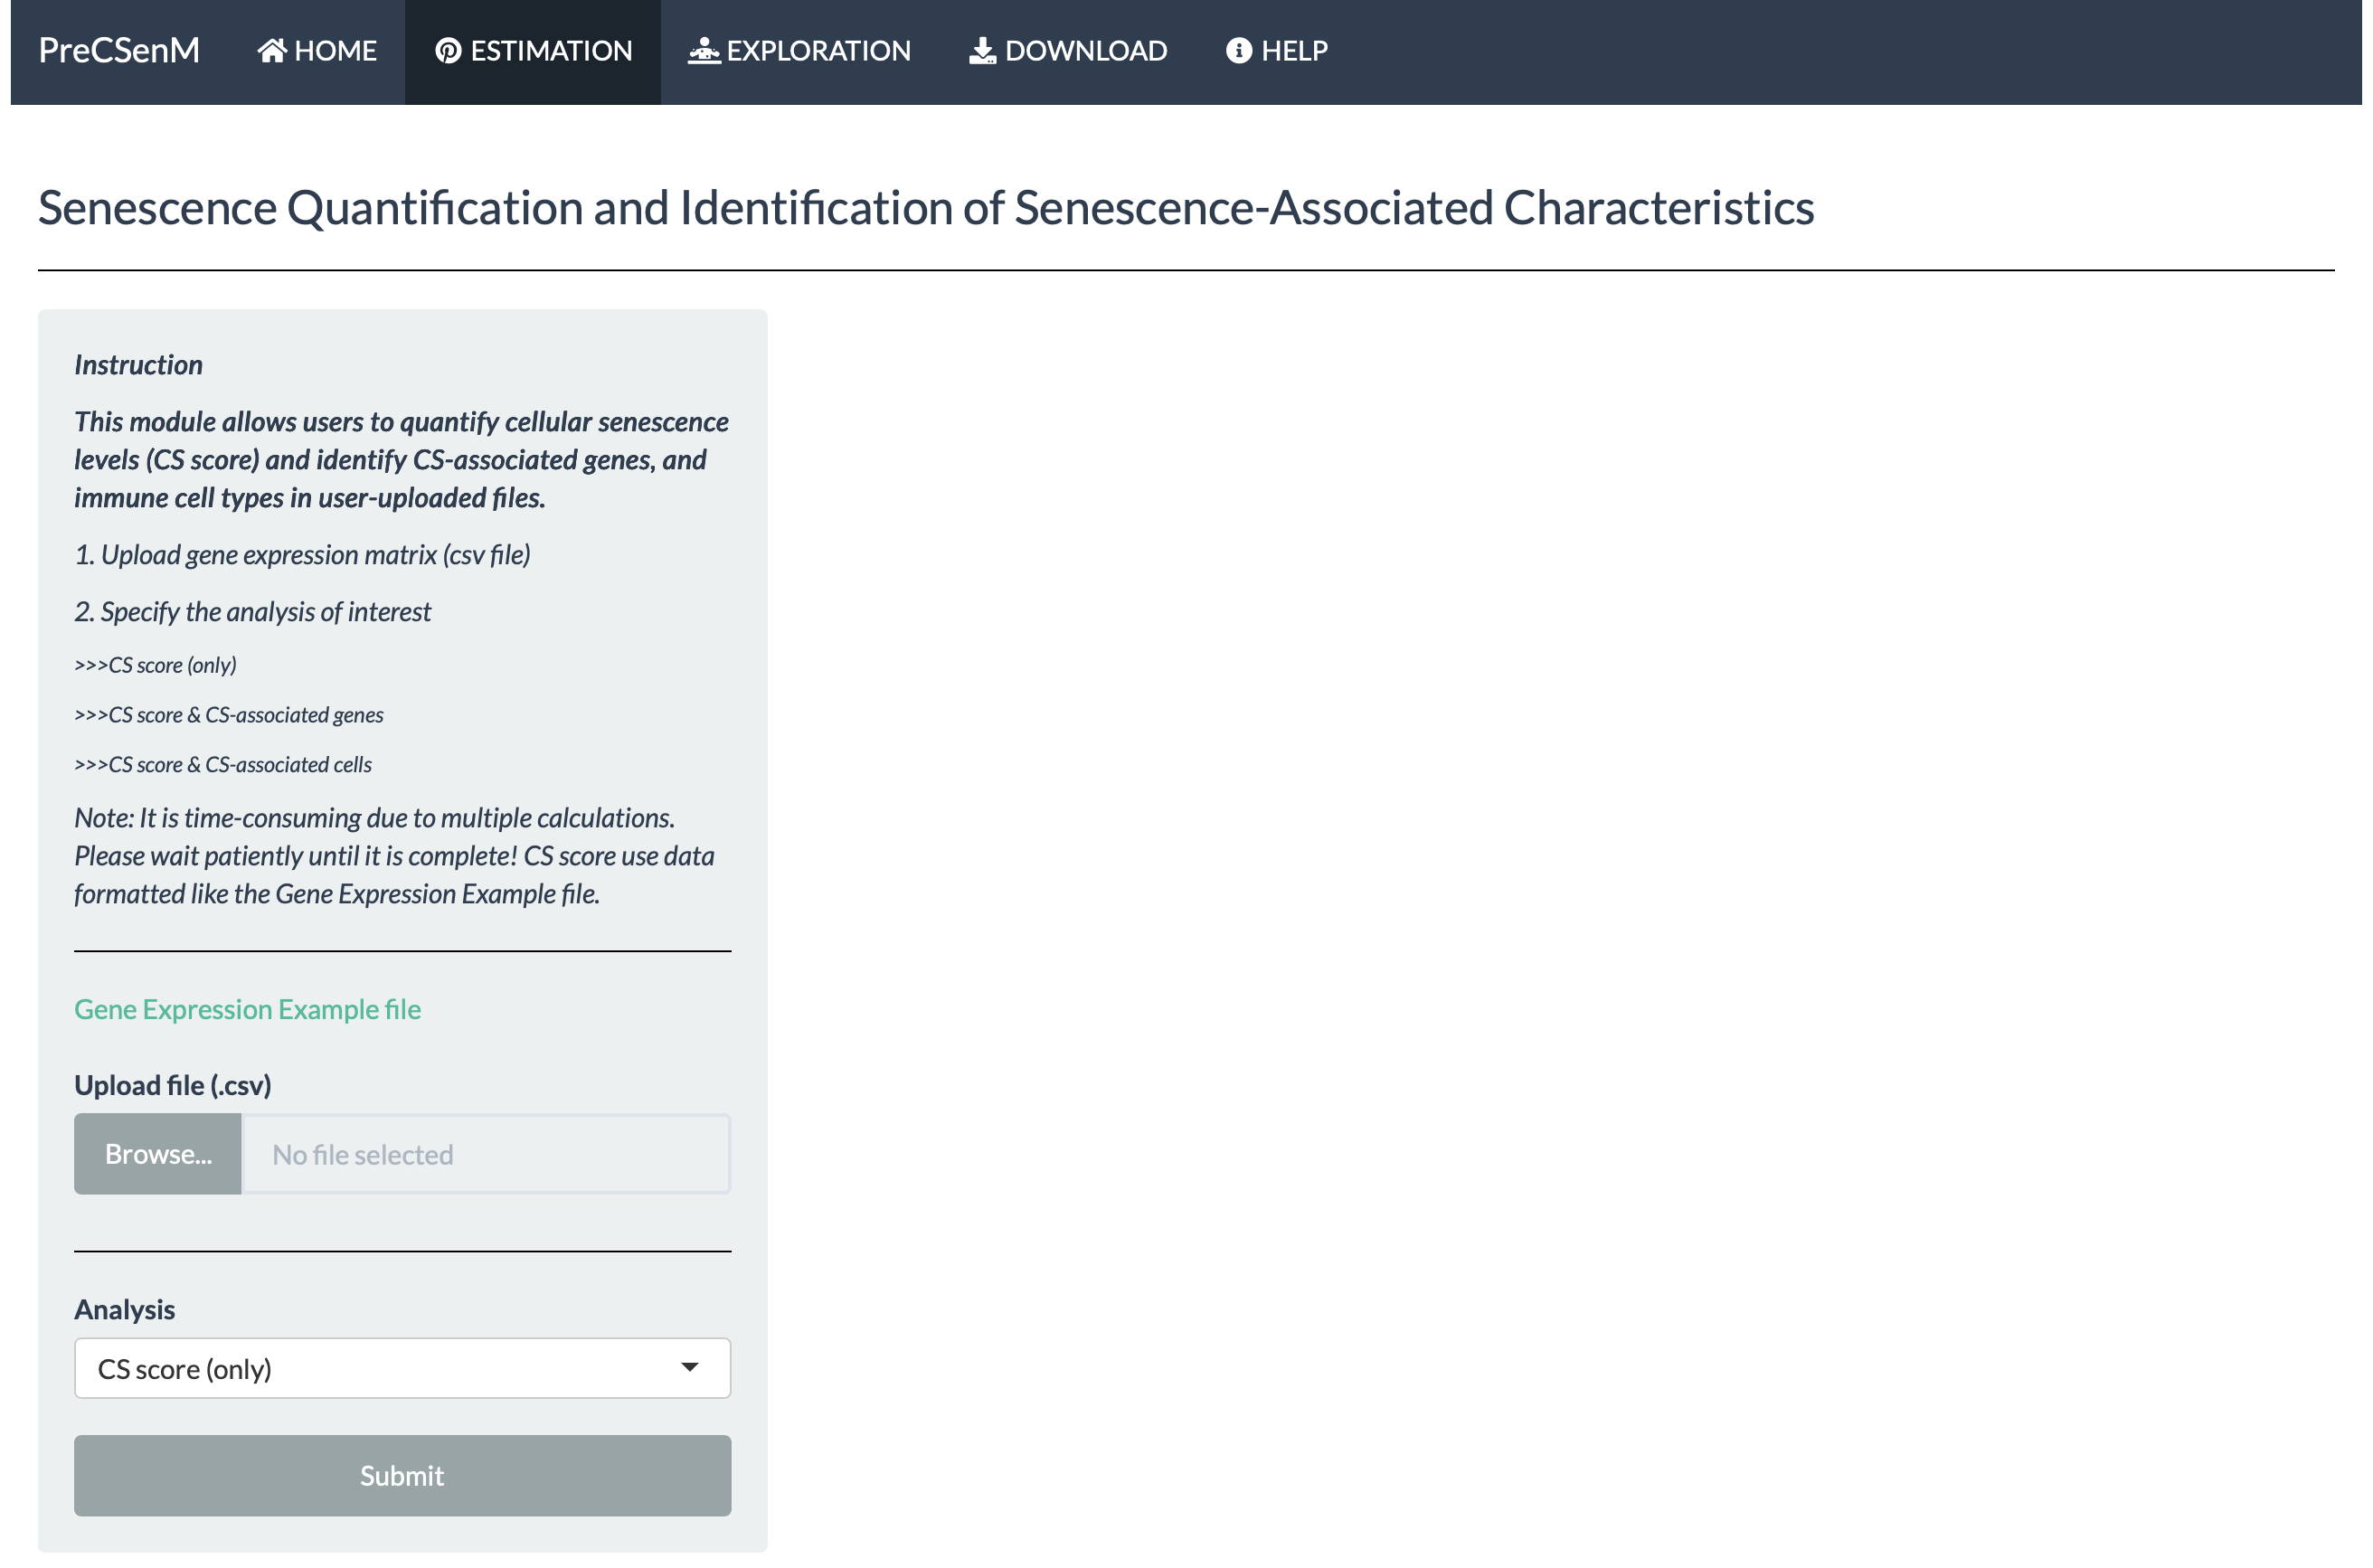
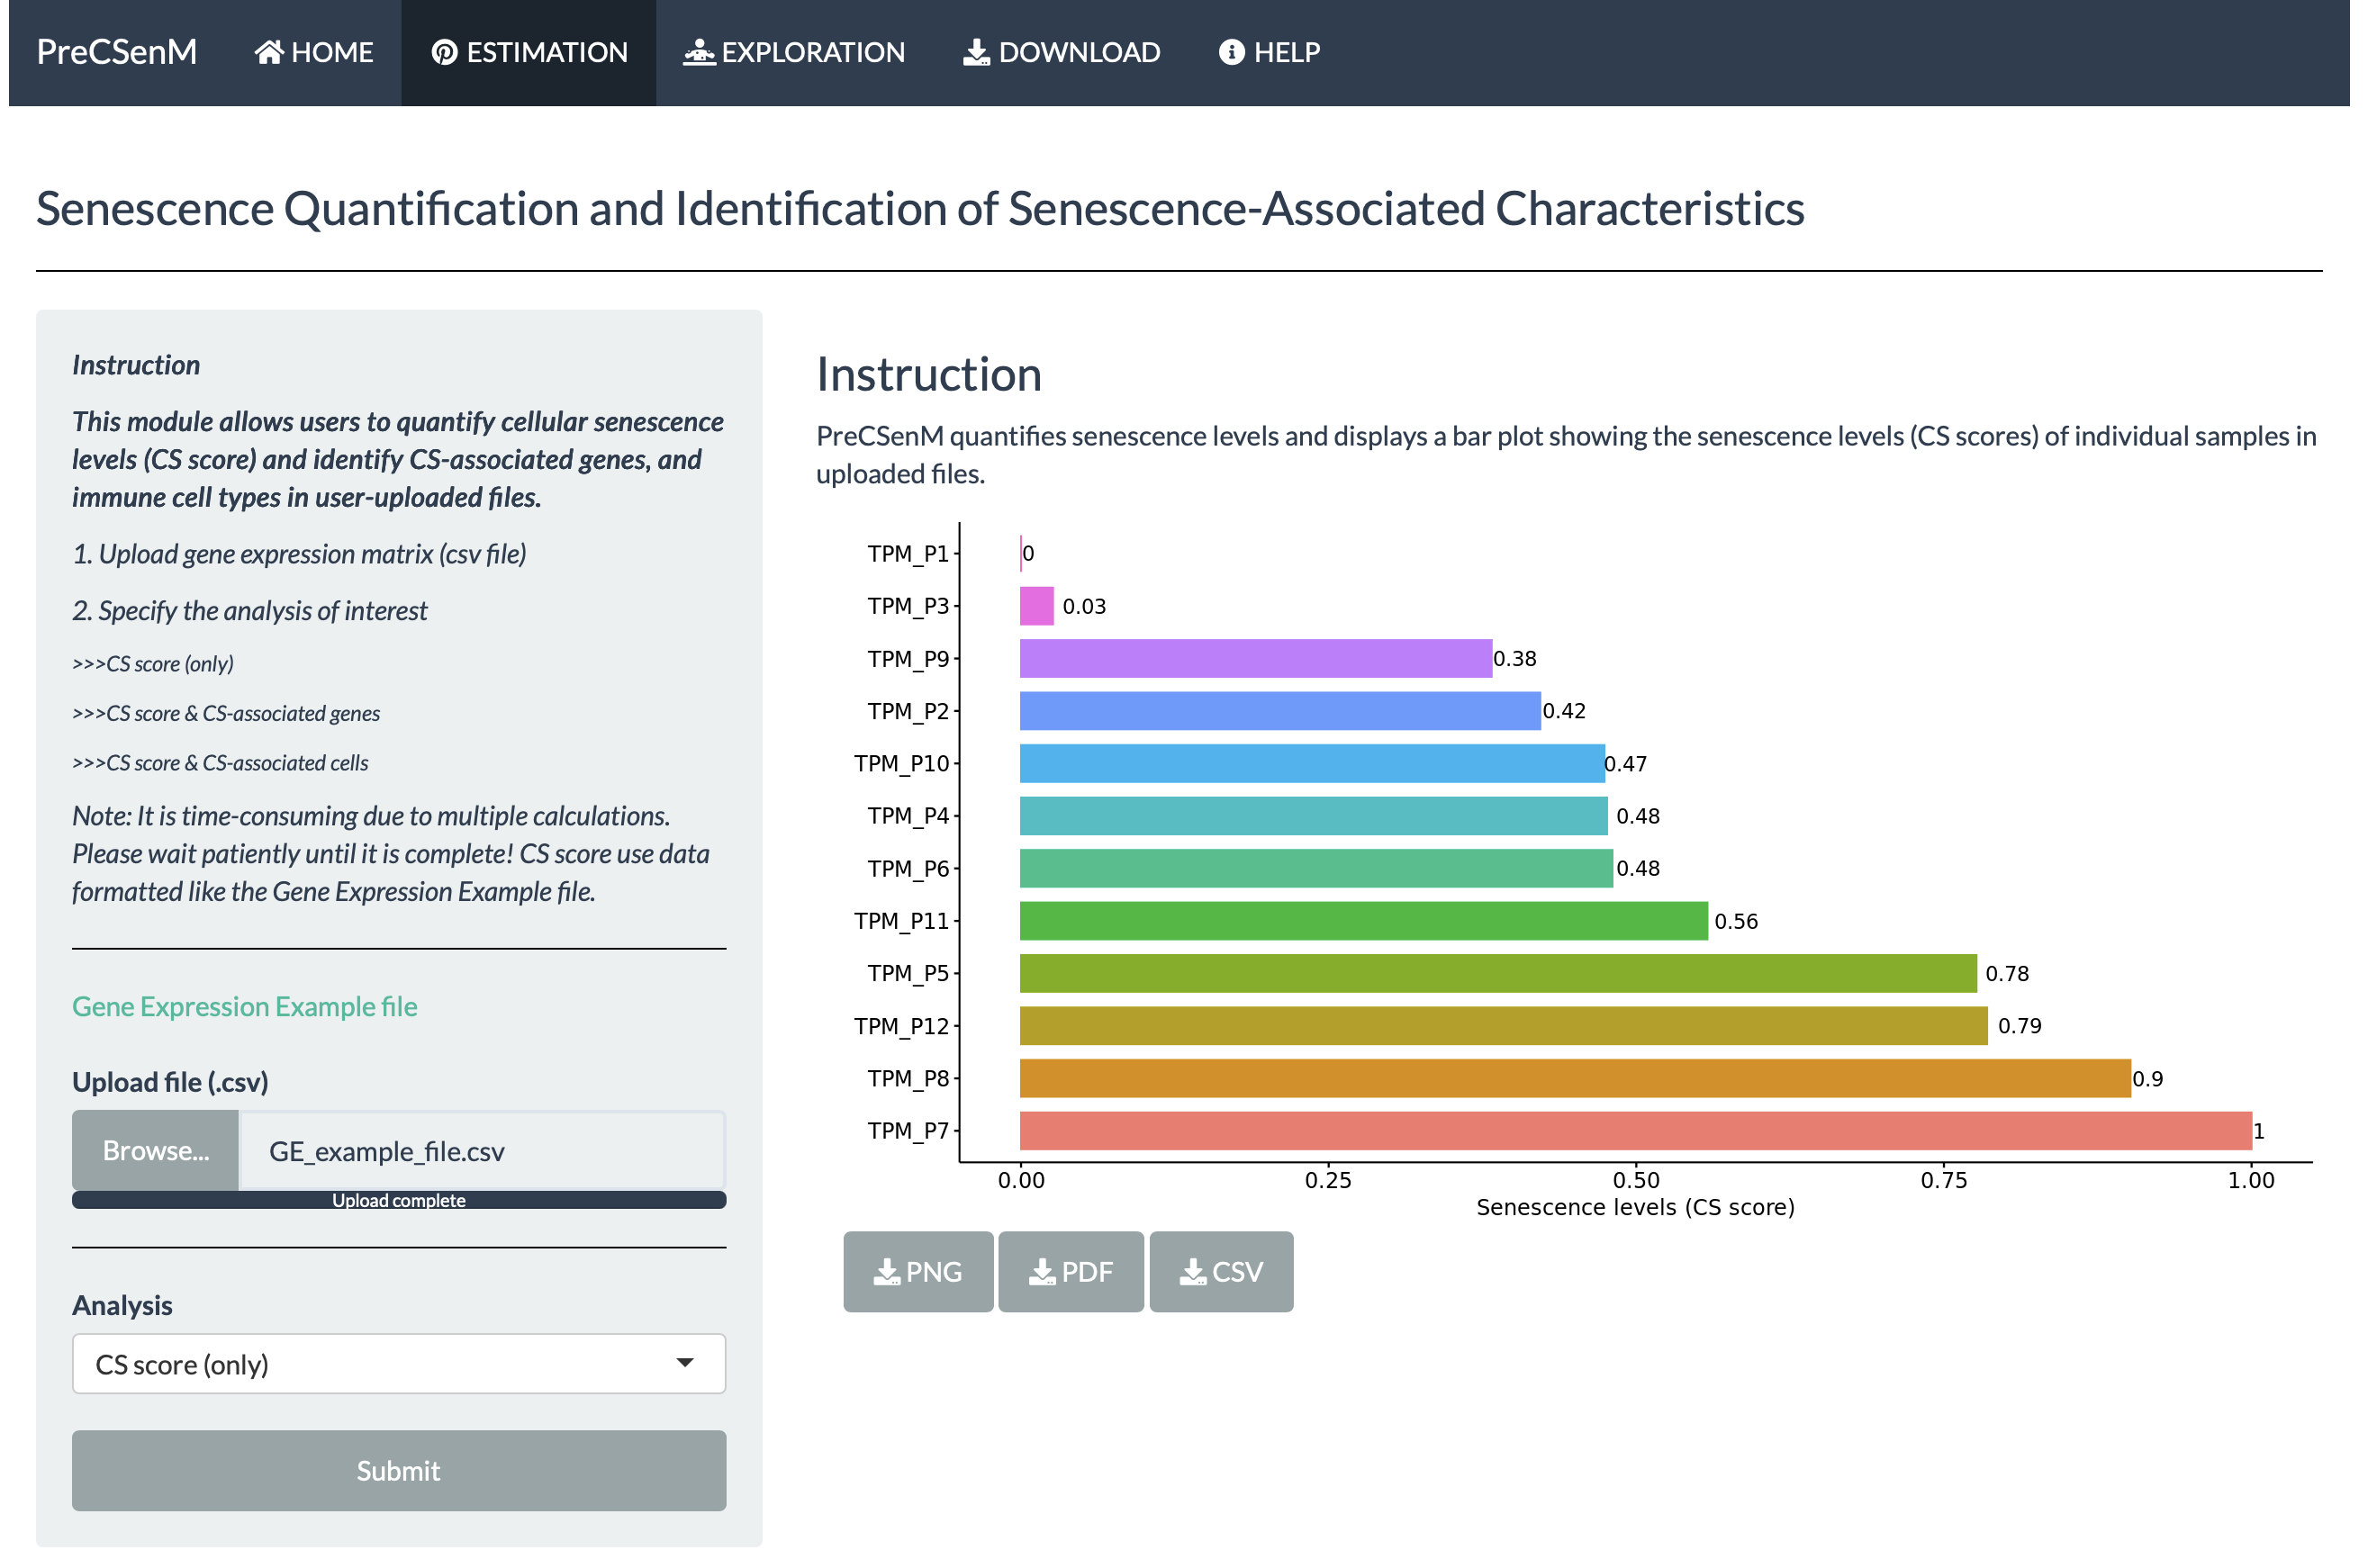
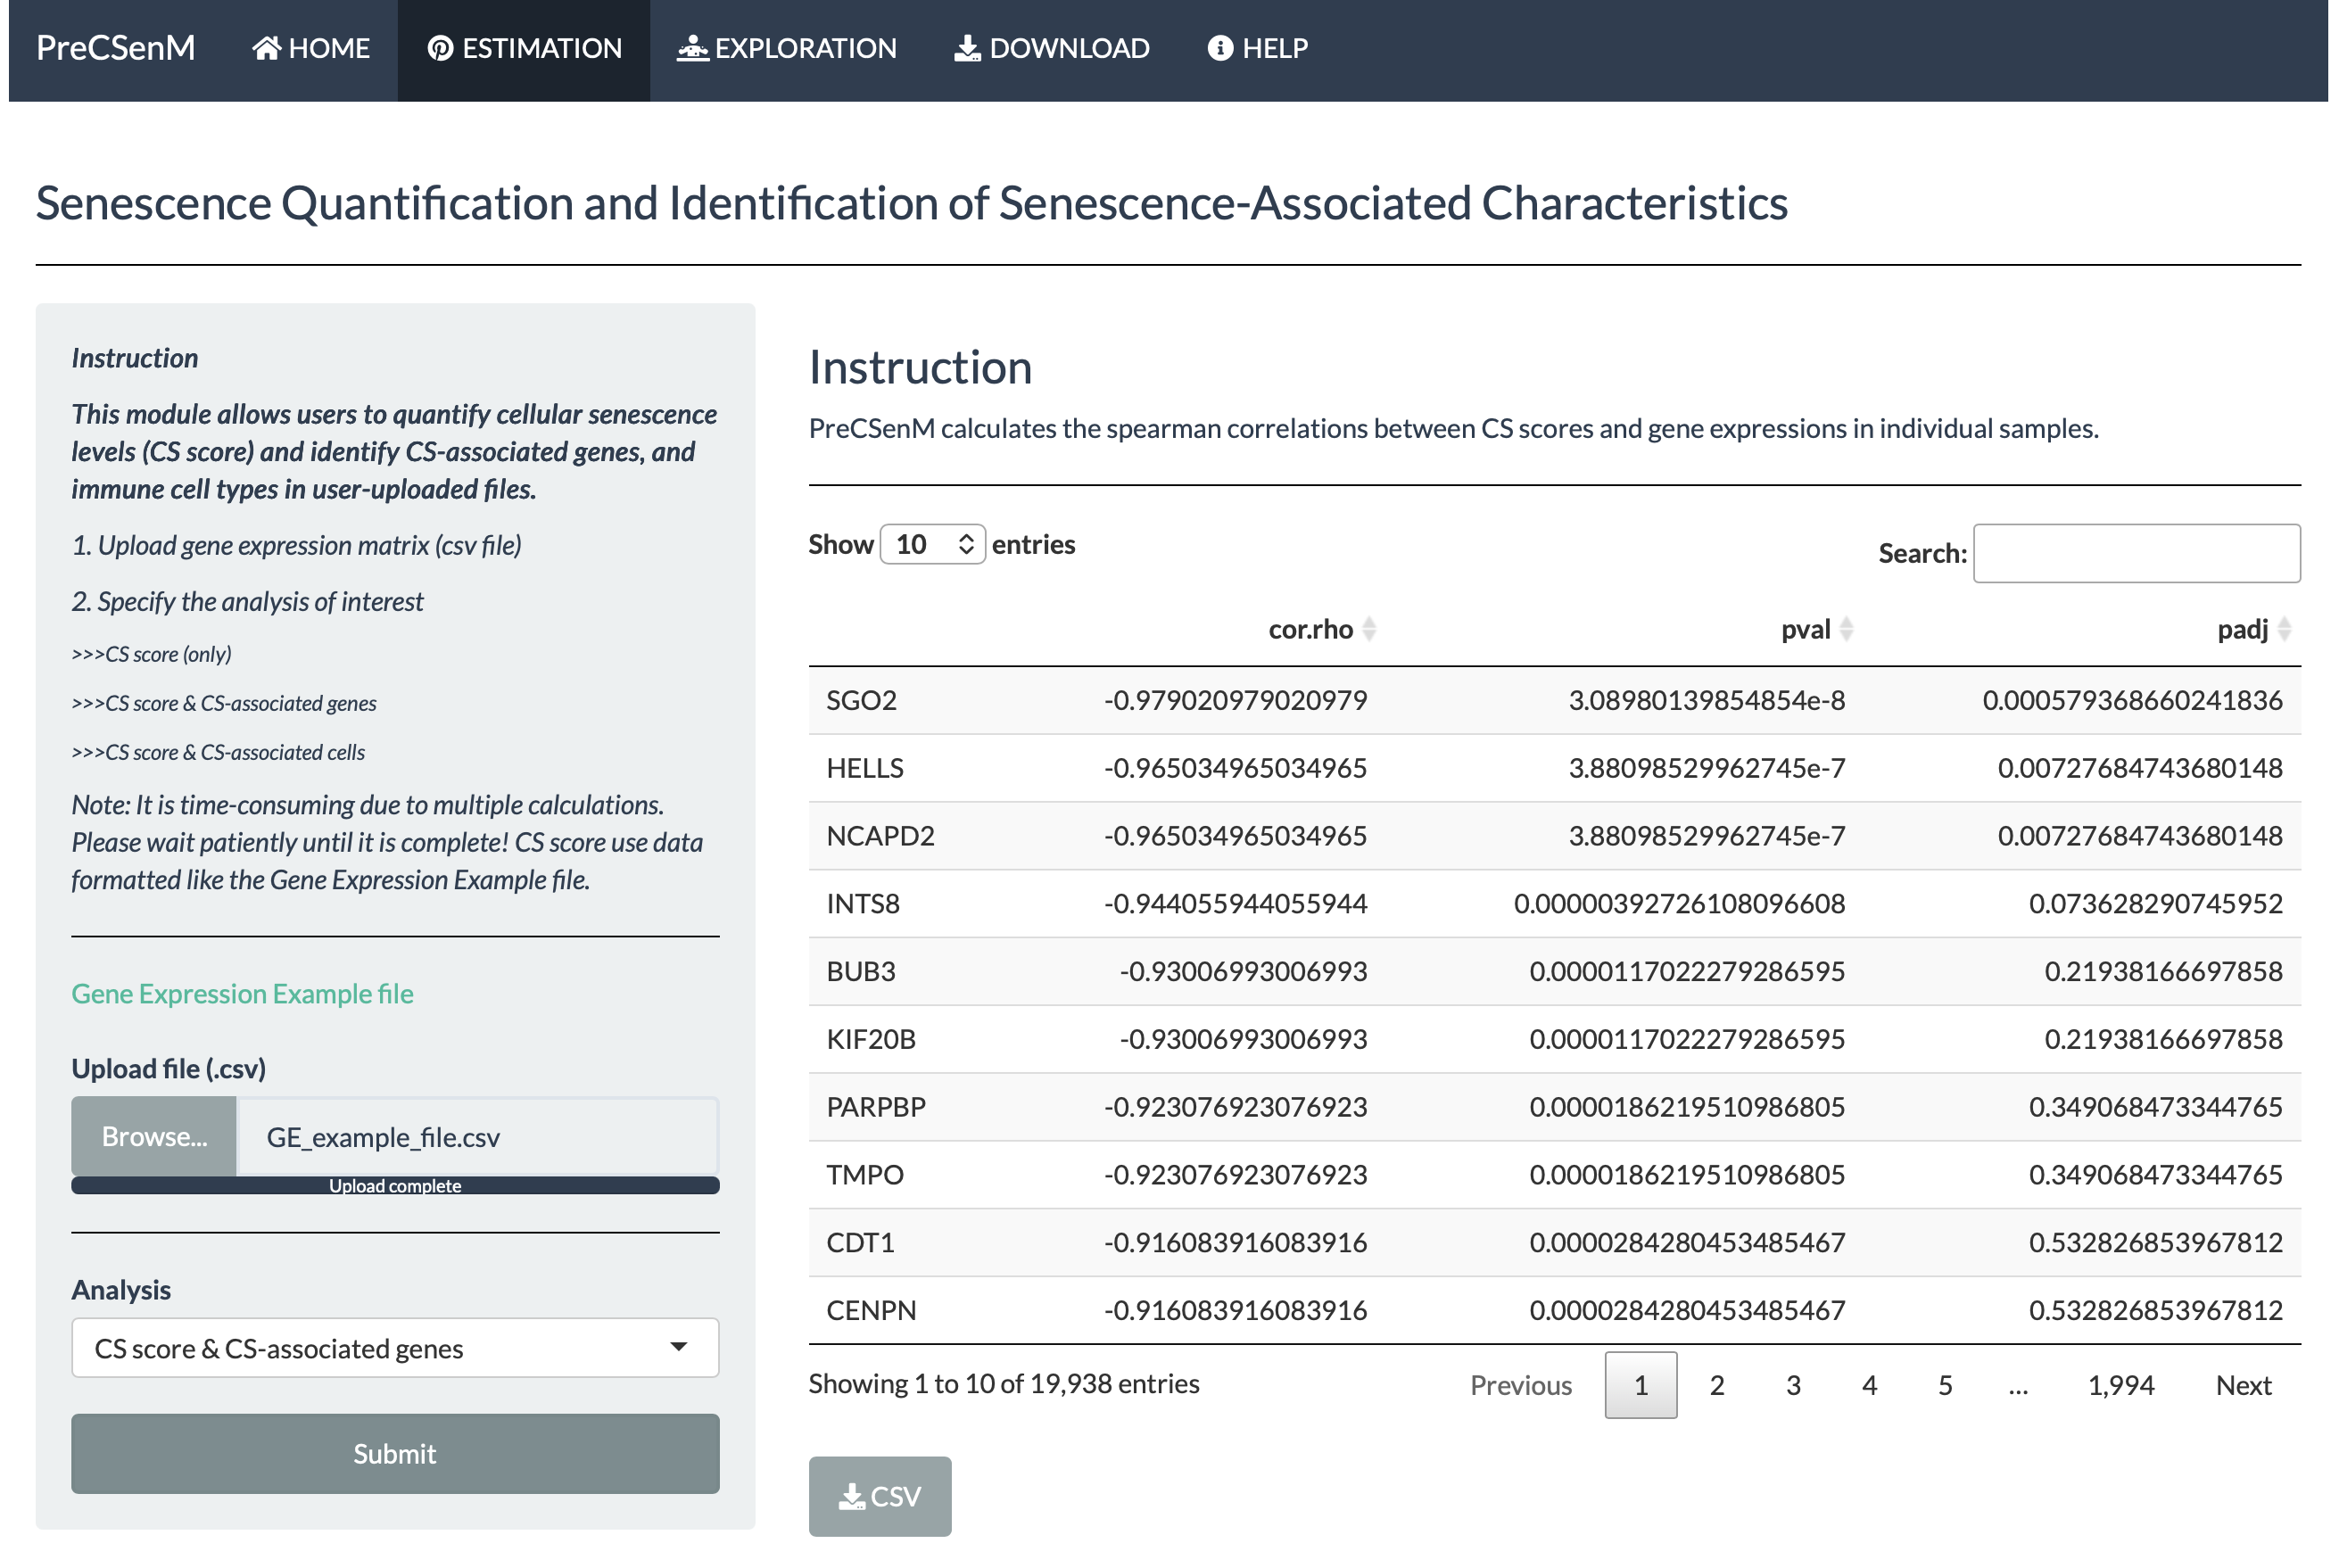
Take example file as an example, select Senescence quantification and Identify senescence-associated genes then click the submit button, a bar plot will be displayed to show the CS scores of individual samples, and the website can identify the correlations between CS scores and gene expressions in individual samples and list CS correlated genes.
PreCSenM also calculates the correlations between CS scores and cell types in individual samples.
Exploration of senescence-associated features in public datasets
Users can explore senescence-associated features in public datasets. The senescence exploration section contains three parts: TCGA analysis, Senescence States Prediction and Single-cell analysis.
*TCGA-LUAD module
In LUAD, our analyses explored the associations between senescence levels and patient survival, genomic alterations, DNA methylation changes, proteogenomic profiles, and pathway enrichment.
*Senescence States Prediction
AUC plot shows prediction of senescent cell line evaluation cohorts (Data_Type,Cell_Type,Induction).
*Single-cell analysis
Users are allowed to select datasets (PRAD_GSE141445, CRC_GSE146771, Glioma_GSE102130, Glioma_GSE131928_10X, Glioma_GSE131928_Smartseq2, NSCLC_GSE117570, NSCLC_GSE143423). UMAP plot shows major cell types and boxplot compares the difference of CS scores across major cell types.
Download
Users are convenient to download files below:
Senescence quantification TCGA.csv.
The description of these two files are listed in the download section.
Cancer type
Cell type in single-cell analysis
Contact us
If you have any questions regarding PreCSenM, please don’t hesitate to reach out to us.
Lifei Ma: lifei_ma AT 126.com; Xiaoman Wang: wangxm815 AT ibms.pumc.edu.cn
Address:
State Key Laboratory of Common Mechanism Research for Major Diseases
Department of Biochemistry and Molecular Biology
Institute of Basic Medical Science
Chinese Academy of Medical Sciences and Peking Union Medical College
Beijing, 100005, China